Pharmacophore-based virtual screening
Find compounds that match your interaction-feature model
Define a three-dimensional interaction-feature model, prepare candidate conformers, rank feature matches, and review a chemically diverse shortlist.
Trusted by 16,500+ researchers and published in
Gather the ligand evidence
A pharmacophore model is built from known actives. Collect actives, and inactives for validation, from ChEMBL or PubChem, and use descriptors to check the sets are comparable.
Prepare 3D conformers
Matching needs 3D structures with sensible hydrogens and charges. Generate and repair them here, then pass them to a pharmacophore program, which ProteinIQ does not provide.
Read the interactions
Features come from how ligands contact a protein. TEMPL scores template-based poses by shape and feature overlap, and ProLIF lists residue contacts. Both help you build a model by hand.
Review the matches
After matching elsewhere, filter the ranked compounds for assay interference and lead-like properties, and add predicted ADMET. Matching a pharmacophore does not show that a compound binds.


Run pharmacophore-based virtual screening on ProteinIQ
Free
$0
For trying and exploring
- Every tool and example result
- 200 credits/month
- 3 jobs per day
- Up to 1,000 residues per job
- SketchIQ, PDB and MSA viewers
- Academic license
Plus
$23 per user per month, billed annually
For academic research
- Everything in Free
- 24,000 credits/user/year
- No daily job limit
- Batch runs
- API and MCP access
- Read-only viewers
Pro
Popular$79 per user per month, billed annually
For commercial work
- Everything in Plus
- 96,000 credits/user/year
- Run workflows
- More parallel jobs
- Larger file storage
- Commercial license
Enterprise
Custom
For large organizations
- Everything in Pro
- Unlimited team members
- Invoice/PO billing
- SAML and SCIM
- Custom model hosting
- Security review
More virtual screening workflows
Structure-based virtual screening
Uses a three-dimensional target to generate and score candidate binding poses, often alongside pocket, property, and pose-quality review.
Shape-based virtual screening
Ranks candidate conformers by three-dimensional overlap with one or more reference ligands, optionally including chemical-feature similarity.
Ligand-based virtual screening
Ranks compounds using similarity, molecular fingerprints, pharmacophores, or learned features derived from known ligands.
Inverse virtual screening
Evaluates one compound against many potential targets to generate hypotheses about intended targets, off-targets, selectivity, or repurposing opportunities.
Read the filters before you trust them
Background guides on the methods, scores, and numbers behind these workflows.
Drugs · 14 min
PAINS filters and pan-assay interference compounds
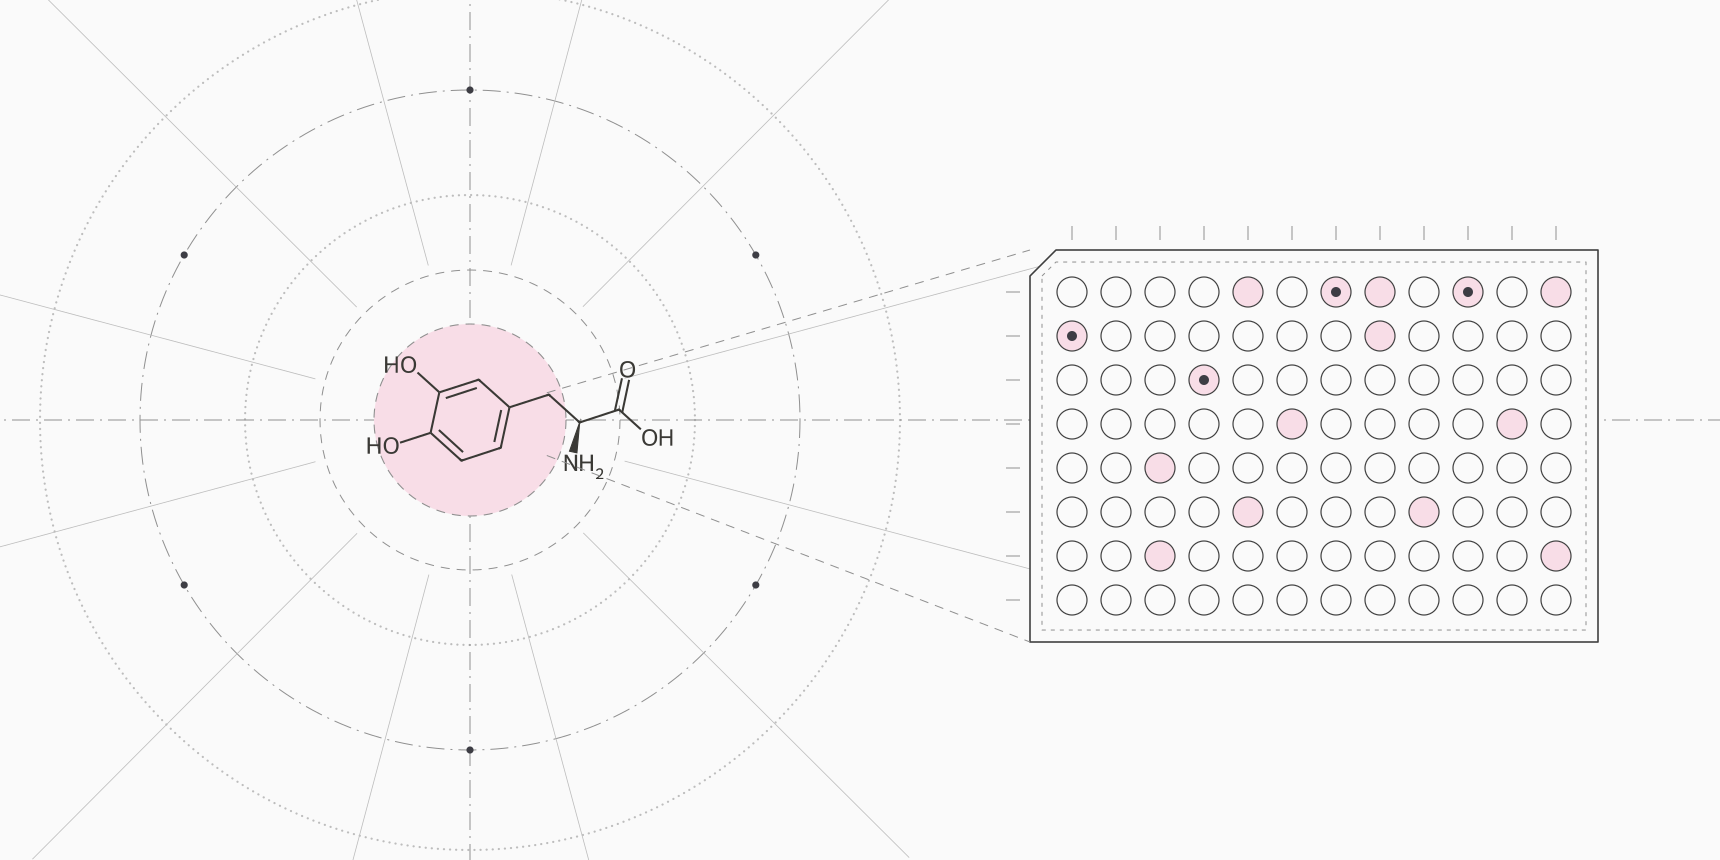
PAINS filters are 480 substructure alerts that Baell and Holloway published in 2010 to flag compounds that often give false hits in screening assays. ProteinIQ ran them on 2,104 approved small-molecule drugs in ChEMBL 37: 122 (5.8%) trigger at least one alert, led by catechols such as levodopa and quinones such as doxorubicin.
Drugs · 14 min
Lipinski's rule of five
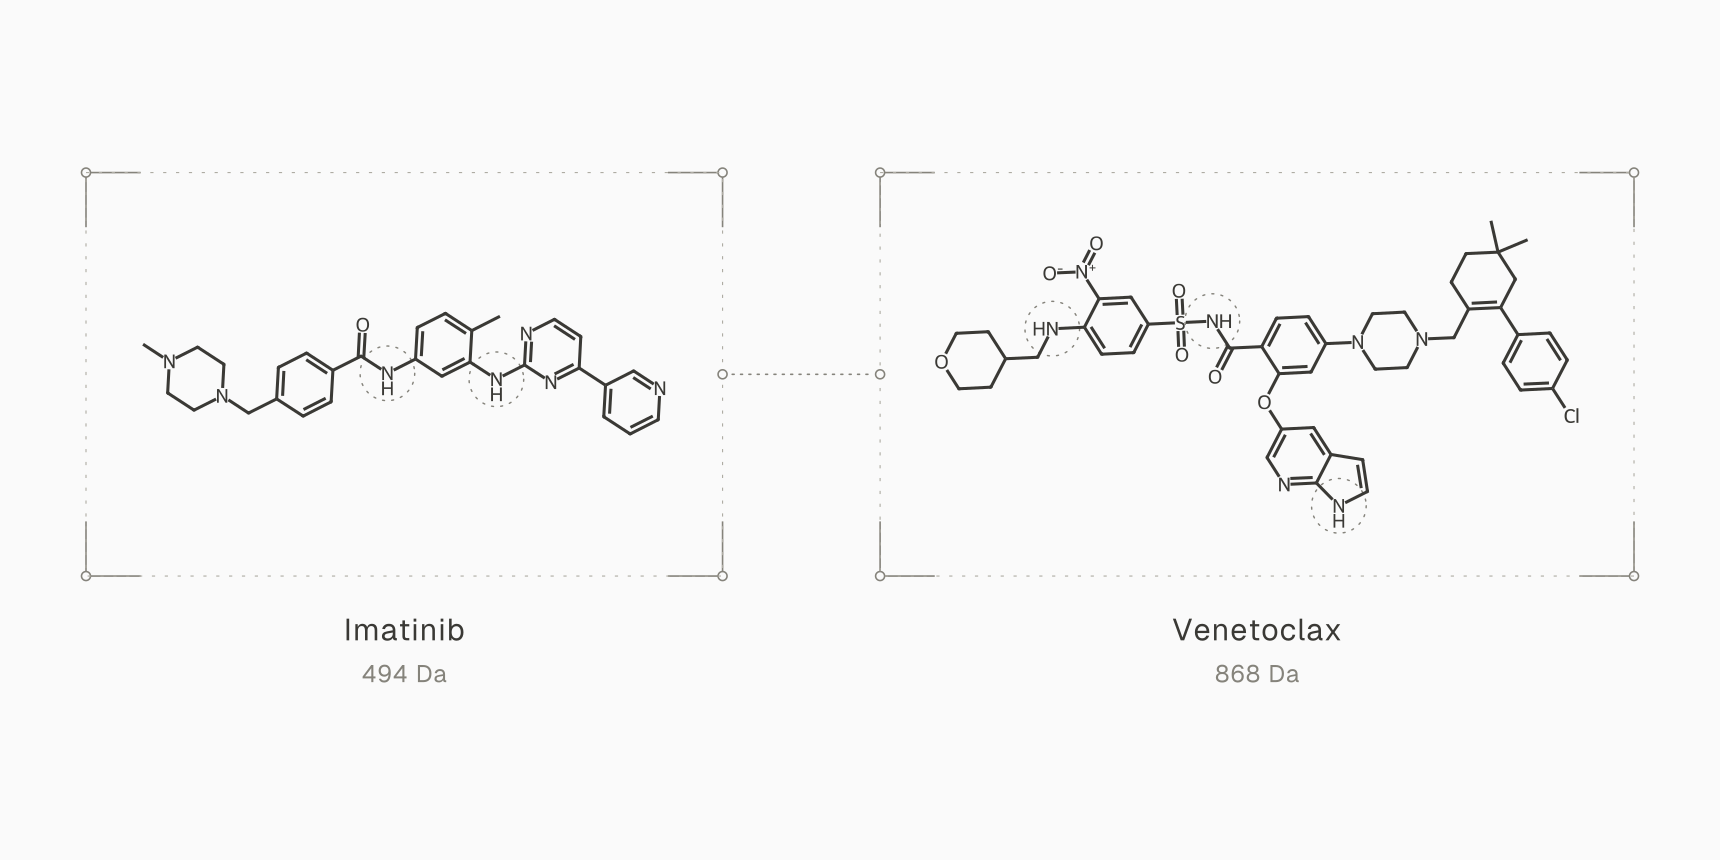
Lipinski's rule of five warns that oral absorption is less likely when a molecule breaks two or more of four limits: molecular weight above 500 Da, cLogP above 5, more than 5 hydrogen-bond donors and more than 10 acceptors. ProteinIQ found that 13% of 2,104 approved small-molecule drugs in ChEMBL 37 break two or more, and 28% break at least one.
FAQs
Use the evidence that is most reliable for the program. Ligand-based models need relevant, diverse actives and credible alignments; structure-based models need suitable complexes and careful treatment of the binding site. Comparing independently derived models can reveal assumptions that one source alone hides.
There is no universal number. Too few features can admit many nonspecific matches, while too many can overfit the known ligands. Choose required and optional features using held-out validation and inspect the chemical diversity of retrieved compounds.
Separate model-building compounds from evaluation compounds. Test recovery of held-out actives against appropriate inactives or decoys, inspect false positives and aligned conformers, and compare more than one plausible model before selecting a production screen.
Not generally. Pharmacophore matching is a feature-geometry filter, while docking samples poses in a target and applies a target-specific scoring function. They can be complementary stages when both the feature model and receptor setup are defensible.
A published 2025 Chemspace offer priced a 1.76-million-compound 3D screen at $5,000 and included shape screening, a pharmacophore-similarity report, analysis, and 200 curated compounds. Because it bundled several deliverables, it is a concrete reference point rather than a pure pharmacophore-only rate.
Current pharmacophore software licenses and contract projects are often quote-based. Model construction, reference or complex curation, conformers per compound, library size, model validation, and manual review usually matter more than a simple per-compound price.
ProteinIQ self-service starts at $29 per month for academic Plus and $99 per month for commercial Pro, with credits quoted before the configured workflow runs. A done-for-you project can be scoped separately around model preparation, screening, and shortlist review.
Review the actual aligned conformer, matched and missed features, stereochemistry, protonation state, conformer plausibility, excluded-volume clashes, molecular properties, and whether the model is applicable to that chemistry before selecting it for follow-up.
AI answers from our docs. No sign-in needed.
Focus on research, not infrastructure
Start free with 200 credits a month and no installs. Upgrade when your project needs more compute.