LocScale
v2.3.1.post5Physics-informed local sharpening for cryo-EM density maps.
Physics-informed local sharpening for cryo-EM density maps.
LocScale performs local density scaling for cryo-EM maps. It sharpens map regions according to local signal and reference information rather than applying one global B-factor across the whole volume.
ProteinIQ runs LocScale on uploaded MRC/MAP files and returns the sharpened map plus the processing log and generated map files. The first release focuses on the standard LocScale command for local sharpening with half-map or full-map inputs. Feature-enhanced maps, coordinate-model refinement, hybrid model completion, MPI runs, and PDF reports are tracked for future support.
| Input | Description |
|---|---|
| Half map 1 | First unsharpened, unfiltered half-map in MRC or MAP format. |
| Half map 2 | Second matching unsharpened, unfiltered half-map. |
| Cryo-EM Map | Full unsharpened map for datasets without half-maps. |
| Mask | Optional MRC/MAP mask. LocScale estimates an FDR mask when this is omitted. |
| Reference map | Optional MRC/MAP model map used directly as the LocScale reference. |
Half-map input is recommended when available because LocScale can use the pair for FSC-based resolution estimation.
ProteinIQ accepts MRC/MAP uploads up to 50 MB per file for this service. This is a ProteinIQ upload limit, not a LocScale file-format limit.
The default settings match LocScale defaults for local sharpening: one CPU process, FDR threshold 0.01, averaging filter size 3, cube size 32, and no input filtering. Optional resolution, window-size, FDR-window, FDR pre-filter, mask-threshold, and reference-map-resolution fields are passed to the corresponding LocScale command-line options only when set.
LocScale returns a primary sharpened MRC map. ProteinIQ also returns the LocScale log and generated processing files such as FDR masks or local-scaling diagnostic maps when LocScale writes them.
Open the returned MRC files in ChimeraX, Coot, or another cryo-EM map viewer for inspection.
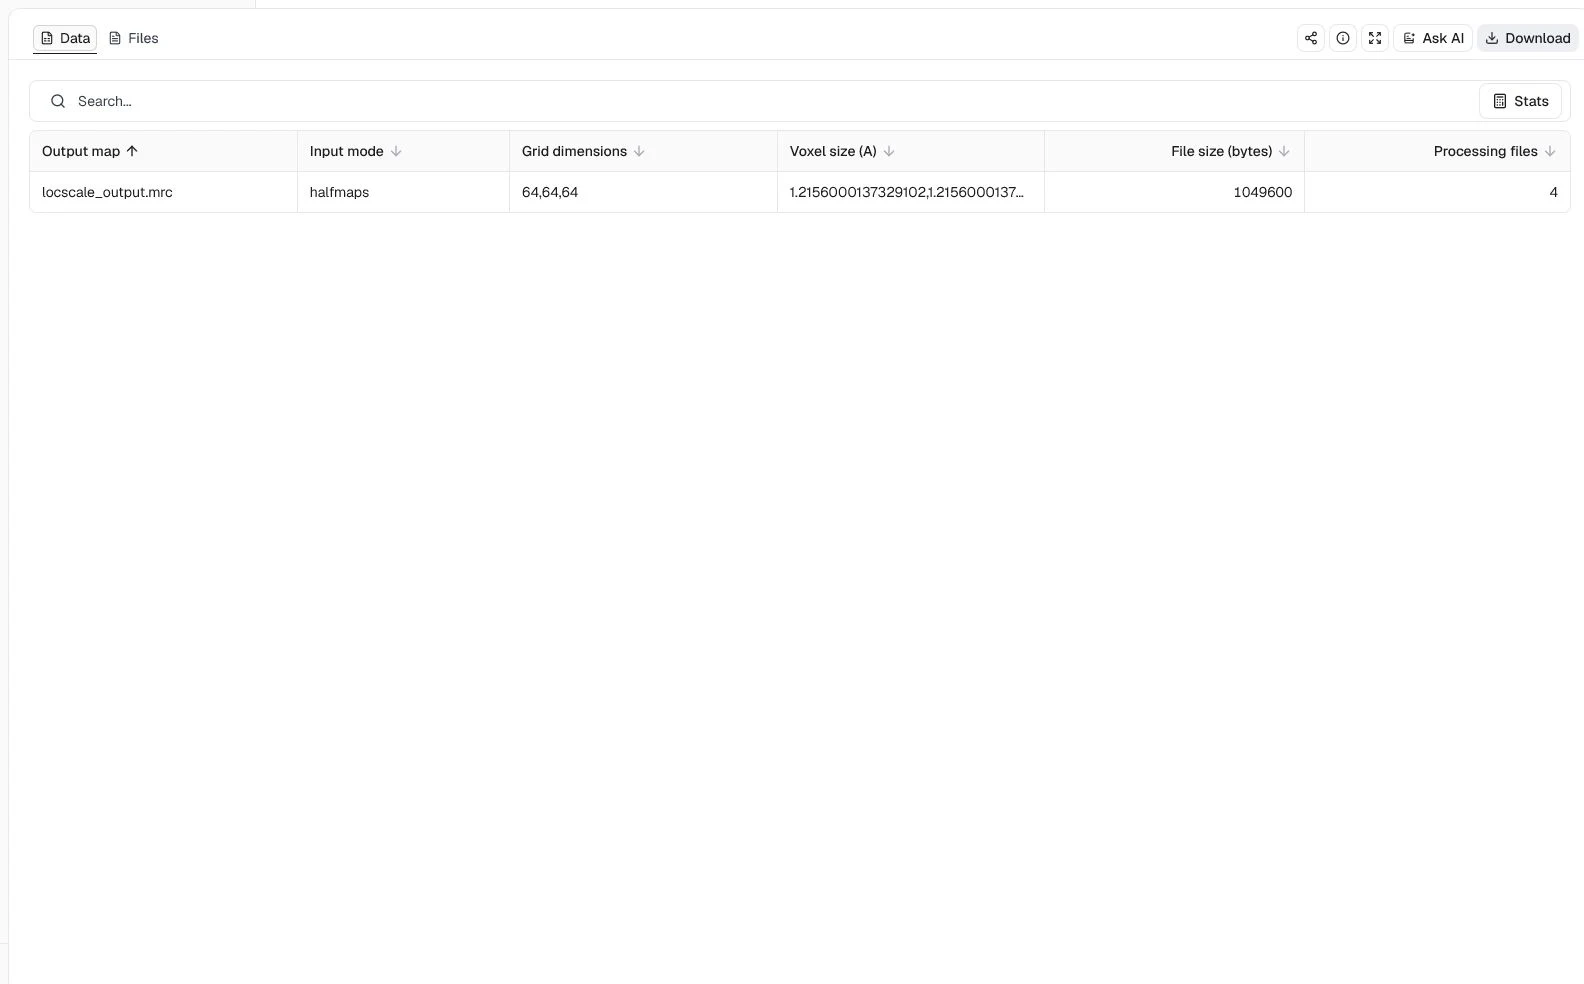
This example demonstrates the recommended paired-half-map workflow on a compact region of the Rattus norvegicus TRPV1 channel dataset EMD-5778. It uses matching unfiltered half-map subvolumes, a data-derived mask, and the corresponding deposited reconstruction as a direct reference map.
CPU processes = 4, Resolution cutoff (A) = 3.4, Window size (pixels) = 24, and Apply FSC filter = enabledThe completed job returns locscale_output.mrc on the original 64 × 64 × 64 grid, together with the reconstructed unfiltered map, bfactor_map.mrc, qfit_map.mrc, and the processing log. The table confirms that LocScale used halfmaps mode and preserved the input voxel spacing. This result shows that the paired-map and FSC-filtering workflow completed; it does not by itself establish improved interpretability. Compare the output with the input half-maps in a cryo-EM viewer before using it for model building.
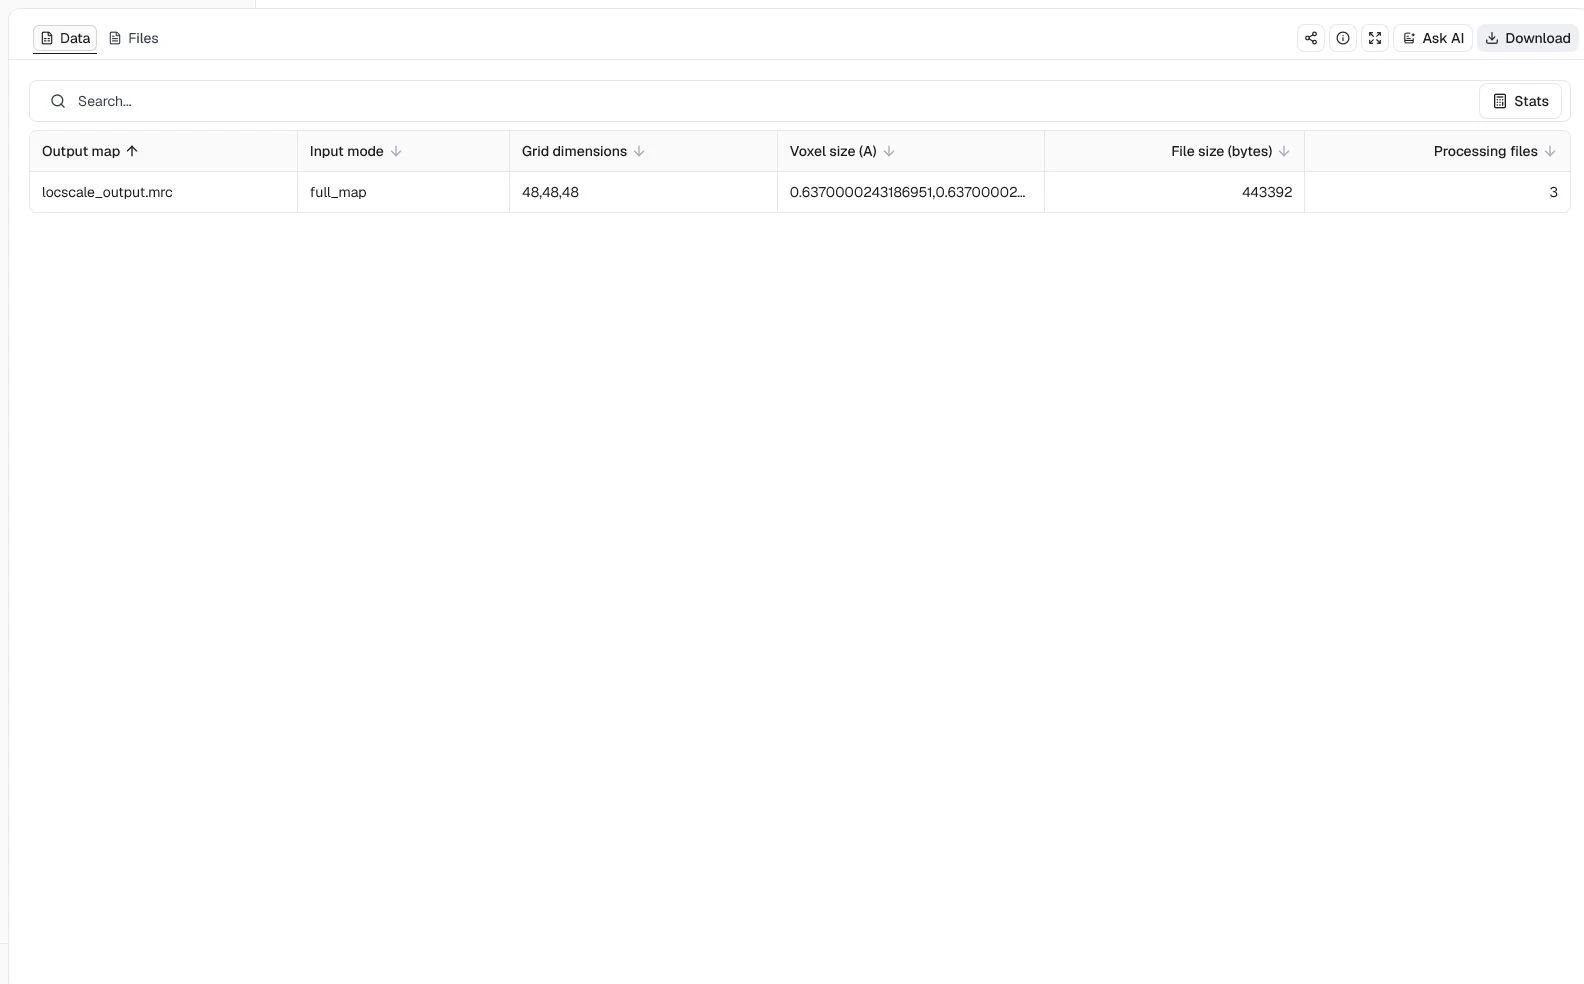
This EMD-2984 example shows full-map operation on a high-resolution region of Escherichia coli β-galactosidase. The input is an average of matching deposited half-map subvolumes, accompanied by a data-derived mask and the corresponding deposited reconstruction as the direct scaling reference.
Map input = Full map, CPU processes = 4, Resolution cutoff (A) = 2.2, and Window size (pixels) = 16ProteinIQ returns a 48 × 48 × 48 sharpened MRC map at the same 0.637 Å voxel spacing, plus bfactor_map.mrc, qfit_map.mrc, and the LocScale log. The compact subvolume makes the reference-map workflow quick to inspect, but it is not a substitute for processing and evaluating the complete EMD-2984 reconstruction.
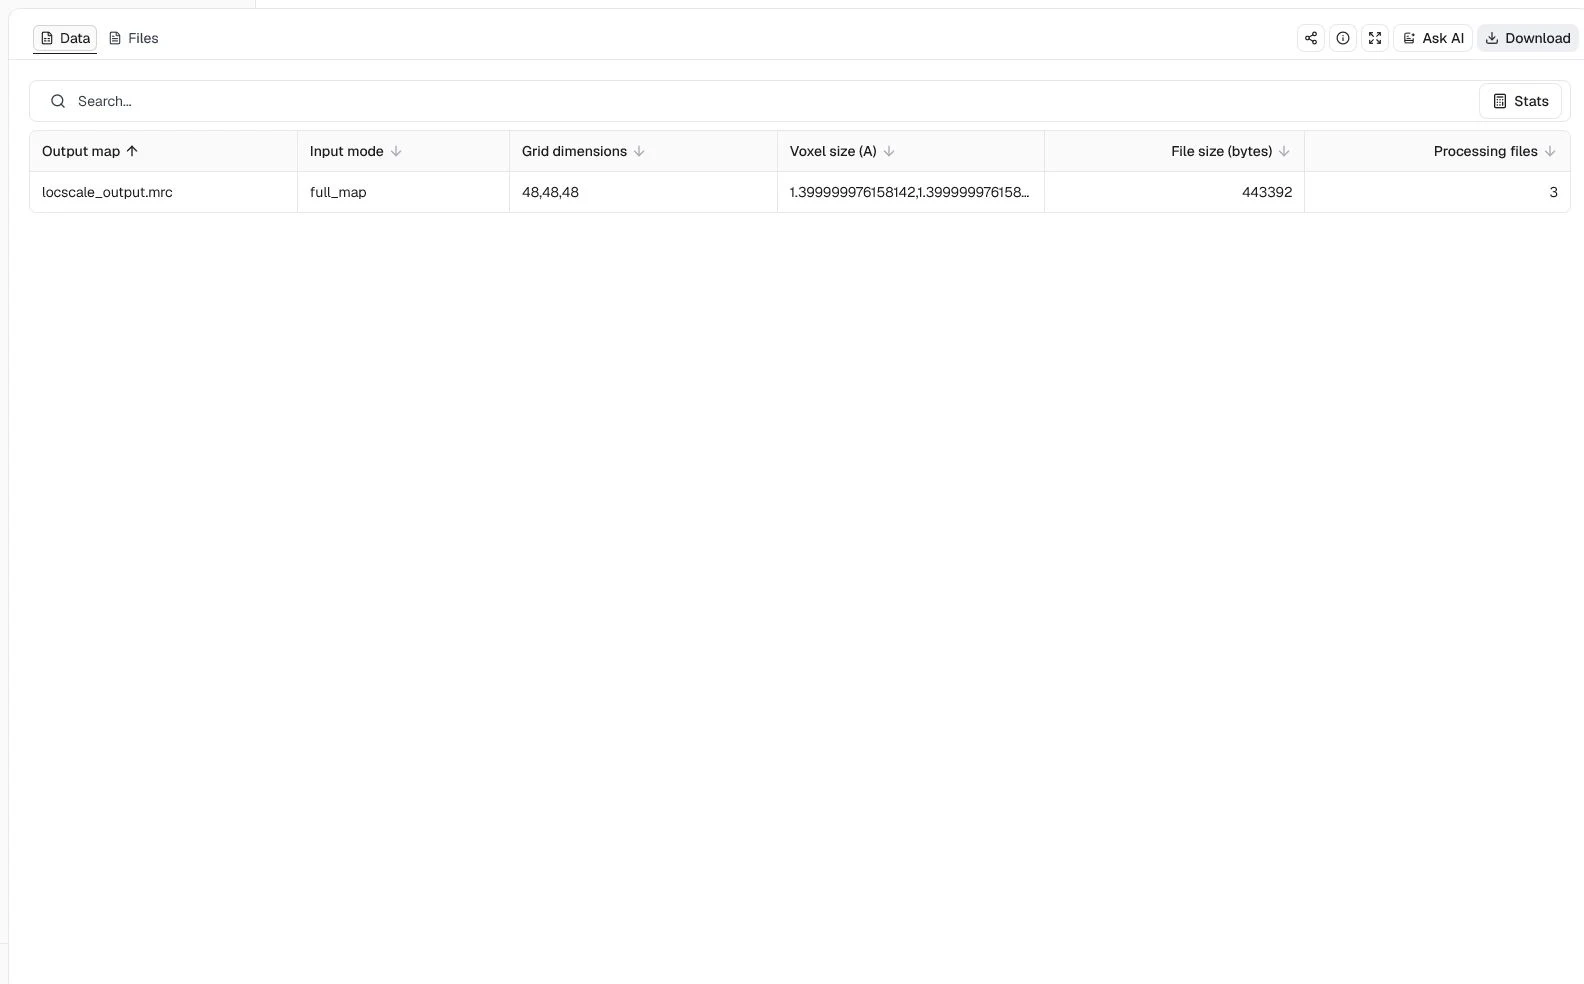
This example uses a compact region from the EMD-3061 human γ-secretase half-map pair to demonstrate full-map processing with both an explicit mask and input filtering. The supplied reference is the matching region from the deposited reconstruction.
Map input = Full map, CPU processes = 4, Resolution cutoff (A) = 3.4, Window size (pixels) = 24, and Filter input maps = enabledThe output table records the expected 48 × 48 × 48 grid, 1.4 Å voxel spacing, and three processing files. This confirms that LocScale completed the masked, filtered full-map workflow and returned its sharpened map and diagnostics. Whether filtering improves local density for a particular structural interpretation still requires direct comparison with the unfiltered input and independent validation against the half-maps.
C1 until symmetry-specific runtime checks are added.
Predict metal and water binding sites in proteins with 3D CNN models.

Validate protein structures with clashscore, Ramachandran, rotamer, and geometry checks.

Generate structural summary reports and diagrams for a single protein PDB structure.

Predict protein-protein binding affinity from structure

Pairwise RMSD matrices for PDB structure ensembles

Diffusion-based hydration site prediction for protein structures

Analyze aggregation-prone regions in a protein structure.

Score every protein interface with ESM-2 and a graph neural network

Assess docking quality between model and native structures

Assign helices, sheets, turns, bends, and PPII structure from atomic coordinates.